Why Pharmaceutical Dissolution Stability Testing Fails: Lessons from Three SAHPRA Recalls

Three products.
Three SAHPRA-approved recalls.
Three dissolution failures that only appeared months after the batches had been released and distributed.

Lasix 80mg failed dissolution at six months. Lumont 10mg failed at twelve months. Cipla-Pioglitazone failed during ongoing stability monitoring. In every case the product passed initial release testing. In every case patients had already received it.

This is the central challenge of pharmaceutical dissolution stability testing: the conditions that cause a product to fail are often invisible at release and only become measurable after months of storage. When the dissolution testing method itself introduces uncontrolled variables, through manual sampling, inconsistent vessel setup or inadequate hydrodynamic control, failures become harder to detect and impossible to defend.

Why do products pass release testing and fail dissolution at stability?

The apparatus controls the dissolution conditions. It does not control what happens to the drug product during storage. That distinction is where most stability failures originate.

Three different storage mechanisms triggered three different failure modes in these recalls. Understanding each one is the starting point for building a dissolution programme sensitive enough to catch them before they become a market withdrawal.
What caused three SAHPRA dissolution stability failures?

Lasix 80mg: furosemide surface passivation

Sanofi Aventis South Africa recalled batch CG022A after dissolution dropped below 80% at 60 minutes at the six-month stability interval. Furosemide is a BCS Class IV drug with dissolution critically dependent on available surface area. During ambient storage, the tablet surface undergoes recrystallisation, converting to a more thermodynamically stable crystalline form that increases hydrophobicity, raises liquid contact angle and progressively retards media penetration. Invisible at release. Measurable at six months.

Lumont 10mg: montelukast excipient matrix hardening

Zydus Healthcare SA recalled Lumont 10mg after out-of-specification dissolution results at the 12-month time point under 30°C and 75% RH long-term conditions. The polymer excipient matrix undergoes cross-linking during extended storage at elevated temperature and humidity. Cross-linking increases structural rigidity, reduces water diffusion rates into the tablet core and creates a denser outer boundary that restricts media penetration. Drug release slows progressively and at month twelve it had slowed below specification.

Cipla-Pioglitazone: particle agglomeration and hydrodynamic dead zones

Pioglitazone is a BCS Class II drug with dissolution dependent on particle size and vessel hydrodynamics. During storage, particles agglomerate, clustering into larger masses that reduce total surface area. In a USP Apparatus 2 vessel those agglomerated masses settle into the near-zero velocity zone beneath the paddle shaft where fluid shear is insufficient to break them apart. Drug release drops. The result is a cone of undissolved material at the vessel base and an out-of-specification stability profile.
 

Why uncontrolled testing variables amplify stability failures

What connects all three recalls is not just the storage mechanism. It is the sensitivity of the dissolution method to detect the developing failure at the earliest possible time point.

Manual sampling introduces timing offsets between vessels. Variable cannula depth changes the concentration gradient at withdrawal. Inconsistent filtration alters the apparent dissolved fraction. Vessel misalignment and imprecise paddle height affect the hydrodynamic pattern within the vessel, including the stagnant zone directly implicated in the pioglitazone failure.

In a formulation where dissolution is already marginal, an uncontrolled testing variable is the difference between catching a failure at six months and missing it entirely until the batch has been on the market for a year.


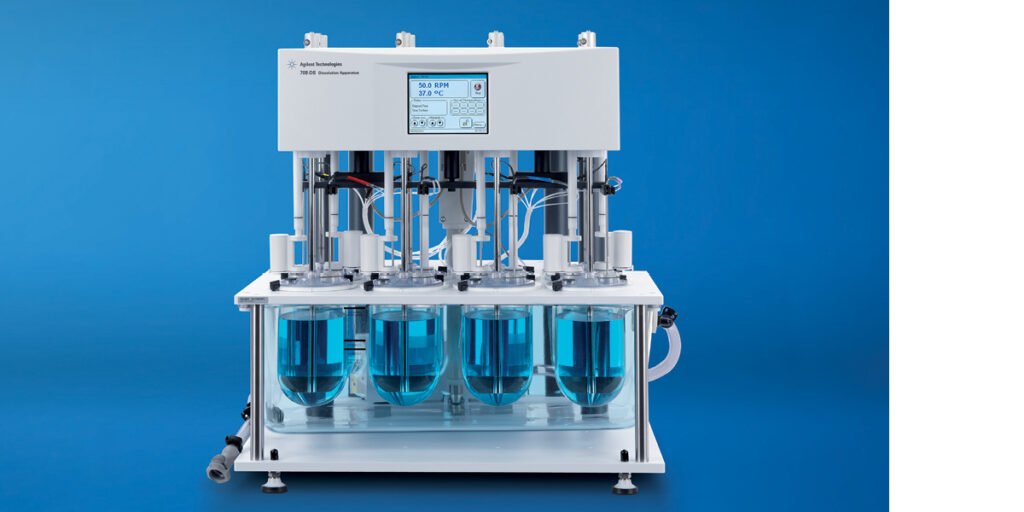
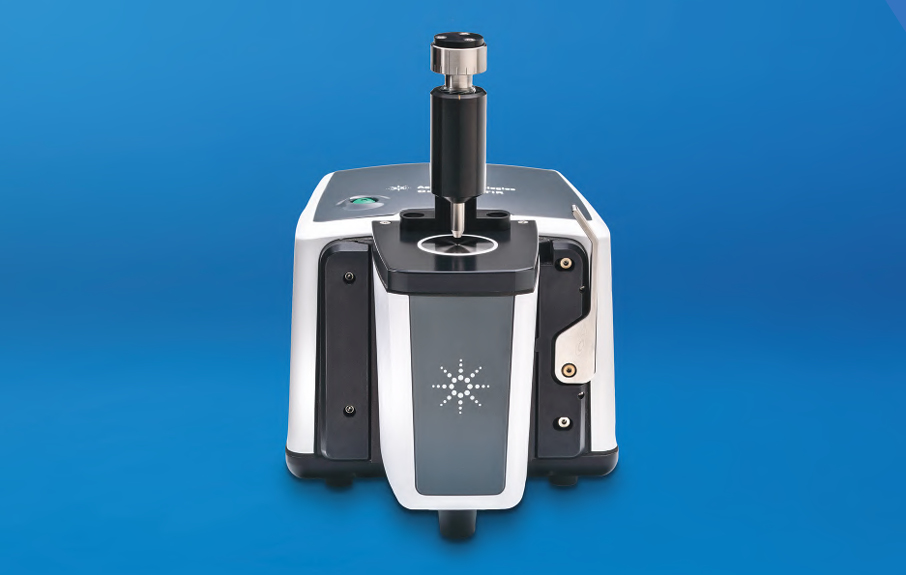
Controlled dissolution testing for pharmaceutical QC: Agilent 708-DS dissolution apparatus

For stability programmes, instrument-level variables must be controlled with the same rigour at month twelve as they are at initial release. The Agilent 708-DS addresses this directly.

TruAlign vessels provide repeatable vessel-to-shaft centering, eliminating the asymmetric hydrodynamic profiles that contribute to stagnant zone formation and cone effect. AutoTemp in-vessel temperature monitoring confirms media temperature is maintained throughout the run. Programmable sampling depth eliminates vessel-to-vessel insertion depth variation. Dissolution Workstation Software provides electronic documentation and audit trail capability for 21 CFR Part 11 compliant environments.
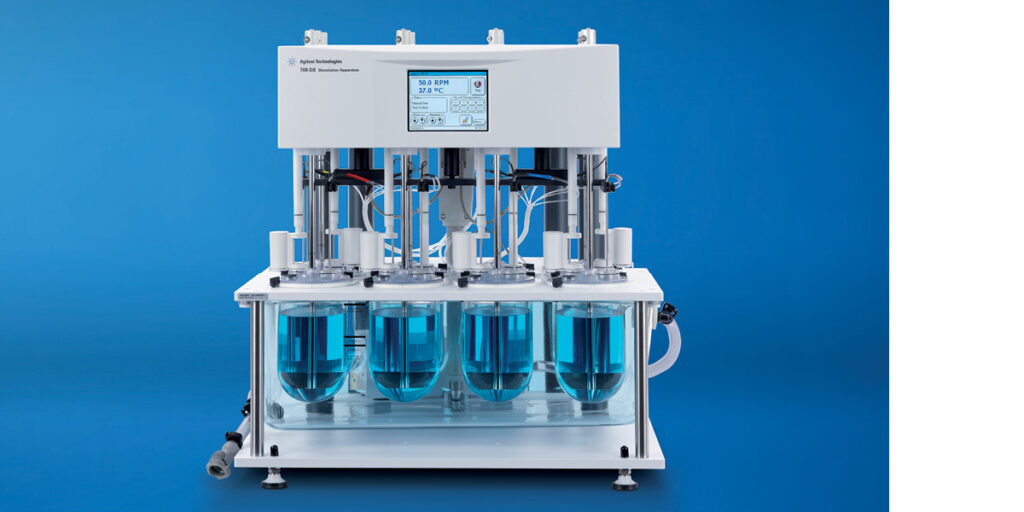
 

Agilent 708-DS

Automated sampling for reproducible stability data: Agilent 850-DS dissolution sampling station

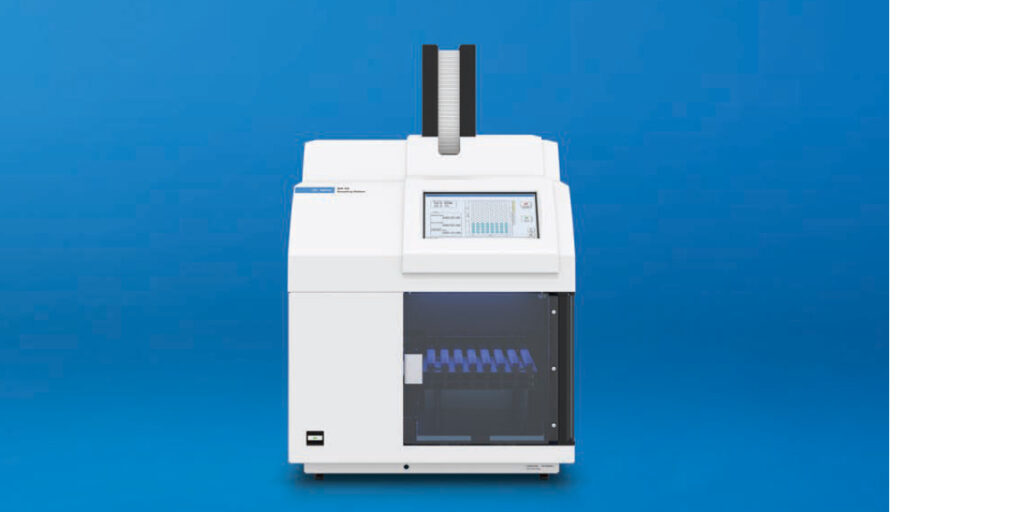
The 850-DS removes the manual variables that amplify dissolution failures in stability testing. Automated withdrawal timing ensures every time point is sampled at the exact programmed interval across all six vessels simultaneously. Automated needle positioning ensures identical sampling depth at every vessel at every time point. Inline filtration at 0.2 or 0.45µm eliminates sample preparation variation from manual filter handling.

For the furosemide, montelukast and pioglitazone dissolution methods specifically, automated sampling is not a convenience. It is a methodological requirement. Detecting a developing stability failure before it becomes a recall requires a testing platform that removes the noise introduced by manual handling.

Practical resource: Agilent 850-DS product page

Speak to a Chemetrix specialist about configuring the 850-DS for your stability sampling volumes, filtration requirements and downstream analytical method.
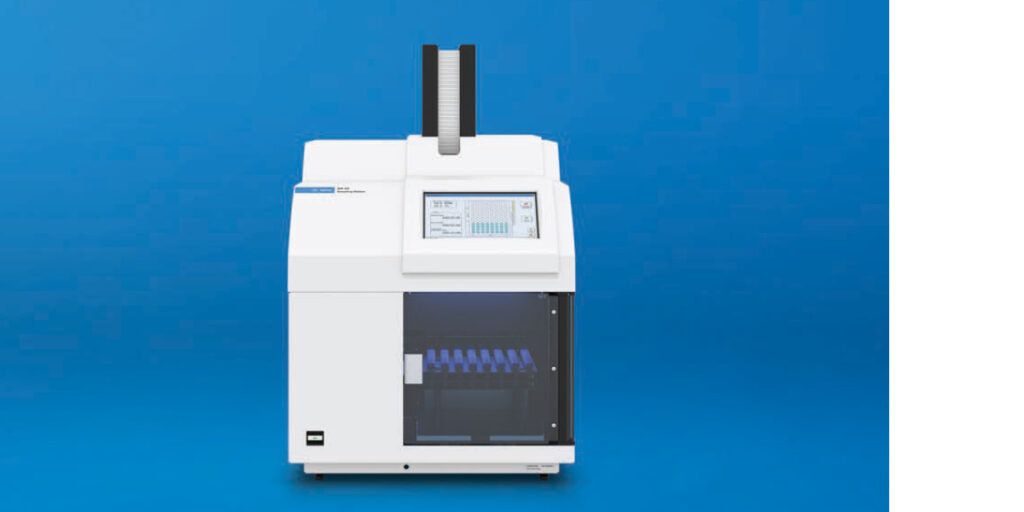
Agilent 850-DS

📥 Explore the full Agilent dissolution testing portfolio
Agilent dissolution testing resources

Passing release testing is the beginning of compliance, not the end of it

Three recalls. Three different storage mechanisms. One shared failure: a dissolution testing methodology not designed to detect the failure early enough to prevent it.

The analysts who run stability dissolution programmes deserve a workflow that is built to detect what needs to be detected; not a manual sampling process that introduces enough variability to mask a developing failure until it’s too late.

Chemetrix works with pharmaceutical QC labs across Southern and East Africa to design dissolution testing programmes built around the sensitivity and reproducibility that stability monitoring actually requires. The goal is dissolution data that defends the product at every stability time point, not just the first one.

Build your stability dissolution programme around the results you need to defend

Before your next stability pull, work through four questions:

  1. Does your method control vessel centering, paddle height and sampling depth reproducibly across analysts and time points?
  2. Are your stability time points sampled within the required interval across all six vessels simultaneously?
  3. Is your filtration consistent enough to distinguish a genuine dissolution change from a methodological artefact?
  4. Does your audit trail support a SAHPRA inspection or GMP compliance review?

Speak to a Chemetrix specialist about your stability dissolution programme:

  • Pharmaceutical QC labs: apparatus qualification, stability method validation and automated sampling configuration for SAHPRA GMP compliance
  • Generic drug development labs: dissolution method development and bioequivalence stability testing for BCS Class II and IV compounds
  • Contract testing labs: high-throughput stability dissolution workflows with HPLC integration and LIMS connectivity
  • Regulatory affairs and QA teams: dissolution data integrity review and method robustness assessment ahead of SAHPRA submissions

Your dissolution data is only as reliable as the stability programme behind it.


TL;DR

Lasix 80mg, Lumont 10mg and Cipla-Pioglitazone all passed initial release testing. All three were recalled from the South African market after dissolution failures at 6 or 12-month stability intervals. The root causes were different in each case but the pattern is the same: subtle physical changes during storage interact with uncontrolled dissolution testing variables to produce results that passing release testing cannot predict. Automated, end-to-end dissolution workflow control is the primary defence.
Pharmaceutical Dissolution Testing: Why Sampling Causes More Variability than the Apparatus

The dissolution apparatus has been validated. Media temperature is correct. Paddle speed is set. Everything looks right. And yet results from one analyst to the next are not as consistent as they should be.

This is one of the most common and least-discussed problems in pharmaceutical dissolution testing. The problem is almost never the apparatus. It is almost always what happens next: the sampling.

Dissolution testing determines how an active pharmaceutical ingredient is released from a dosage form under defined conditions. A failed or inconsistent result can delay batch release, trigger a data investigation and consume analyst time that could be spent elsewhere. When that variability traces back to sampling rather than the product itself it is a workflow problem, not a formulation problem. And it’s a solvable one.

Why does pharmaceutical dissolution testing produce variable results even when the method
is correct?

The apparatus controls the dissolution conditions. It does not control what happens when the sample leaves the vessel. That distinction is where most labs have a gap.

Manual sampling introduces variability at five distinct points: when the sample is taken, where in the vessel it is withdrawn from, how much is collected, how it is filtered and transferred, and how consistently that process is repeated across multiple vessels and time points. In extended-release products requiring sampling over several hours the opportunity for cumulative variation is significant.

The result is a dissolution profile that reflects the sampling process as much as the drug product itself. Chemetrix consultants see this pattern consistently across pharmaceutical QC labs: unexplained batch-to-batch variation traced back not to formulation but to manual sampling steps that were never formally controlled.

Dissolution testing explained

Every oral dosage form that reaches a patient has been through dissolution testing. The test simulates conditions the drug encounters in the body, exposing the finished product to a defined volume of media at controlled temperature and stirring speed, then measuring API release at defined time points.

Those results determine whether a batch is released for patient use. They underpin bioequivalence decisions in generic drug development and support regulatory submissions across USP, EP and JP pharmacopoeial frameworks. They are used throughout the product life cycle from R&D through to routine QC.

When dissolution data is compromised by sampling variability the consequences extend beyond a failed batch. They reach into data integrity queries, audit findings and regulatory submissions built on data that does not accurately reflect the product. The analytical lab’s responsibility is to produce dissolution data that reflects the drug, not the process of collecting the sample.


How can pharmaceutical QC labs reduce analyst-to-analyst variation in dissolution sampling?

Remove the manual steps. Not because analysts are careless but because repetitive manual sampling across six vessels at five or more time points is an inherently variable process regardless of analyst experience.

The labs that achieve the most consistent dissolution data are not necessarily the ones with the most skilled analysts. They are the ones that have replaced analyst-dependent sampling variables with automated controls. When timing, withdrawal position and filtration are managed by the sampling station the analyst is freed to review data, prepare subsequent runs and apply scientific judgement where it actually matters.

This is what Chemetrix means by treating dissolution as an end-to-end workflow. The apparatus handles dissolution conditions. The sampling station handles the rest. Both need to be
designed together.

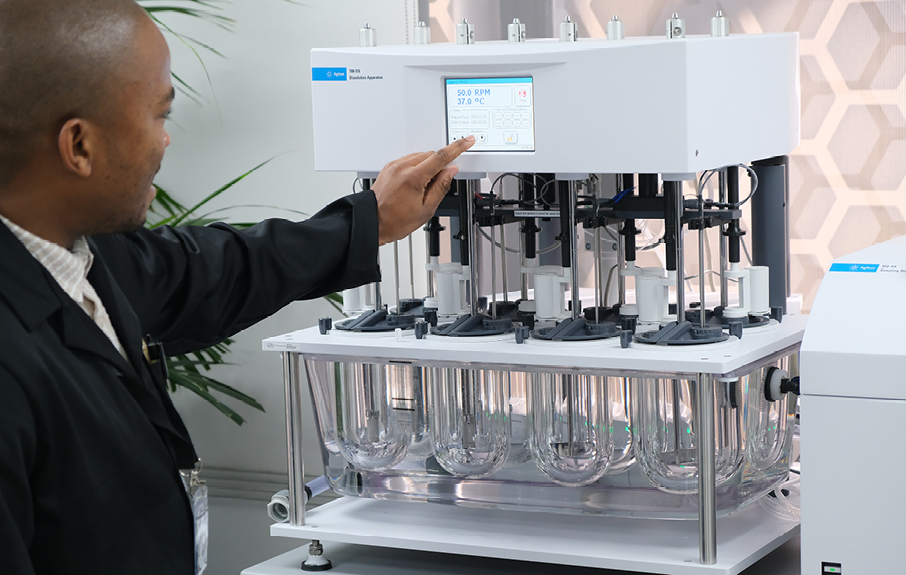
Controlled dissolution testing for pharmaceutical QC: Agilent 708-DS dissolution apparatus

The Agilent 708-DS is the established platform for pharmaceutical dissolution testing across oral solid dosage forms, topicals, implants, transdermal patches and medical devices. It conforms to international pharmacopoeial specifications for Apparatus 1, 2, 5 and 6 and accommodates tablets, capsules, transdermal and semisolid formulations.

Key features that directly address variability at the apparatus level include TruAlign vessels for repeatable vessel orientation, motorised lift for integration with automated sampling systems, AutoTemp in-vessel temperature monitoring to confirm media conditions throughout the run and programmable sampling depth to eliminate vessel-to-vessel inconsistency in withdrawal position. The optional Dissolution Workstation Software provides electronic documentation of test parameters and audit trail capability for 21 CFR Part 11 compliant environments.

For labs requiring online UV analysis, the 708-DS pairs with the Agilent Cary 60 UV-Vis Spectrophotometer in multicell and fiber optic configurations enabling real-time dissolution profile monitoring without manual sample withdrawal at each time point.

Speak to a Chemetrix specialist about configuring the 708-DS for your specific dosage form and regulatory requirements.

Agilent 708-DS

Automated dissolution sampling for regulated pharmaceutical labs: Agilent 850-DS dissolution sampling station

The Agilent 850-DS Dissolution Sampling Station automates what the apparatus cannot control. It manages withdrawal timing, sampling position, filtration and sample collection into test tubes, HPLC vials or 96-well plates depending on the downstream analysis method.

For labs running HPLC analysis, samples collected by the 850-DS into Agilent HPLC trays can be placed directly into the HPLC autosampler without manual vial transfer. A validated end-to-end workflow demonstrated with multidrug capsules containing four APIs at different concentration ranges showed that the full sequence from dissolution sampling through LIMS integration to HPLC measurement and result reporting can be completed with minimal manual intervention. Barcoded vials scanned into the LIMS were automatically linked to vessel and time point data, passed to Agilent Sample Scheduler software and submitted to the LC instrument without the analyst managing the sequence manually.

The 850-DS includes an integrated syringe pump, optional filtration module for 0.2 or 0.45 µm filtration, variable pump speeds for different media types including surfactants, automated cleaning cycle and user access level controls to prevent unauthorised method changes.

Speak to a Chemetrix specialist about configuring the 850-DS for your sampling volume, filtration requirements and downstream analytical method.

Agilent 850-DS

📥 Explore the full Agilent dissolution testing portfolio

Dissolution data quality is a workflow decision, not just an instrument decision

There is a mindset Chemetrix consultants encounter regularly: that some variability between analysts or time points is an acceptable part of dissolution testing. That the apparatus is doing its job and the rest is just the nature of manual work.

It is not acceptable. Not when the data produced determines whether a product reaches a patient. The analysts running dissolution tests day after day under time pressure and throughput demands deserve a workflow that sets them up to produce reliable data consistently, not a process that makes accuracy dependent on replicating the previous analyst’s technique.

The labs that consistently produce defensible dissolution data treat sampling as a controlled variable from method design stage. They invest in the workflow that connects the apparatus to the result. Chemetrix works with pharmaceutical QC labs to design and supply that complete workflow, from instrument selection and qualification through to sampling automation and data management. The goal is dissolution data you can defend, not variability you have learned to explain.

How to diagnose dissolution sampling variability in your lab

Before assuming the problem is the product, work through four questions:
  1. Where is variability entering your sampling process? Timing, position, volume or filtration?
  2. Are your sampling time points being hit consistently across all vessels?
  3. How many manual transfer steps exist between the dissolution vessel and your analytical instrument?
  4. Does your current documentation trail support a 21 CFR Part 11 or GMP audit?
Speak to a Chemetrix specialist about your dissolution workflow:
  • Pharmaceutical QC labs: apparatus qualification, automated sampling configuration and Dissolution Workstation Software for 21 CFR Part 11 compliance
  • Contract research and testing labs: high-throughput dissolution workflows with HPLC integration and LIMS connectivity
  • Generic drug development labs: bioequivalence dissolution method development and semi-automated sampling for extended-release formulations
  • Compounding and hospital pharmacy labs: small-volume dissolution testing and method validation support for non-standard dosage forms

TL;DR

Most pharmaceutical labs assume that once the dissolution apparatus is correctly set up the test is essentially controlled. In practice sampling is where the majority of variability enters the workflow. Timing errors, inconsistent withdrawal depth and manual filtration steps all introduce analyst-to-analyst variation that the apparatus itself cannot prevent. The Agilent 708-DS Dissolution Apparatus combined with the 850-DS Dissolution Sampling Station automates these critical steps to produce more consistent reproducible dissolution data with significantly less manual intervention.div

Up the Stack: Why the Real Frontier of AI Lives Between Your Laboratory Instruments

A single high-resolution instrument run can generate several gigabytes of data in minutes. That sounds like progress, and it is, until that data sits isolated in a proprietary format that nothing else in the lab can read without manual intervention. This is the quiet ceiling that modern laboratories are hitting: the first wave of “smart” instrument features, automated peak integration, predictive maintenance alerts, solved real problems inside individual instruments, but they never solved the problem between them.

For lab directors, IT architects, and operations managers, this matters because the next phase of laboratory digital transformation isn’t about smarter instruments. It’s about whether the software stack above those instruments can finally talk to all of them at once.

Why do “smart” lab features stop solving problems at the instrument level?

Automated peak integration and predictive maintenance alerts were a genuine step forward; they reduced manual review and caught failures before they became downtime. But Chemetrix’s consultants have seen the same pattern repeat across regional labs: once those features are deployed, the bottleneck doesn’t disappear. It moves. The constraint shifts from “is this instrument performing well” to “can this instrument’s data talk to the next system in the workflow.”

A chromatography system might integrate its own peaks flawlessly and still hand off a result that a LIMS, an ELN, or a second instrument can’t parse without a manual export-and-reformat step. The smart feature did its job. The workflow around it didn’t get any smarter.

Why does fragmented lab data matter beyond the bench?

The instruments driving modern science, high-resolution mass spectrometers, multi-detector chromatography systems, generate data at a scale that simply didn’t exist a decade ago. A single run can produce several gigabytes of fragmented output, structured around a specific vendor’s proprietary format rather than the workflow the data is meant to serve.

This isn’t just an IT inconvenience. Every manual reformatting step is a point where data can be transcribed incorrectly, where context gets lost, and where audit trails become harder to defend. In regulated GxP environments, this kind of fragmentation isn’t a productivity issue, it’s a compliance exposure waiting to surface during an audit.

How does declarative orchestration differ from standard lab integration?

Most current lab integration is imperative: an IT team manually maps Instrument A’s output format to Instrument B’s expected input, instrument by instrument, vendor by vendor. It works, but it’s rigid. Every new instrument, every software update, every vendor change requires re-mapping the connection by hand.

Declarative orchestration works differently. An analyst sets a scientific goal, “run this sample through this workflow”, and the software layer coordinates the instrument fleet to execute it, regardless of which vendor built which piece of hardware. This shift is only possible when the data underneath is described using open, vendor-neutral standards rather than proprietary formats that lock data inside a single system.

This is where frameworks from the Allotrope Foundation, the Pistoia Alliance, and SiLA become directly relevant. The Allotrope Foundation promotes industry standards that enhance data interoperability, aligning laboratory data practices with FAIR and ALCOA+ principles to support data integrity and compliance. SiLA promotes AnIML, a basic-level data format that counters the problem of lab instruments speaking different languages, aiming to align communication between manufacturers using a shared XML-based data format. Together, these standards are what let agentic AI move from a chatbot overlay sitting on top of a single system to genuine cross-instrument orchestration

Closing the data gap for pharmaceutical and GxP labs

For regulated pharmaceutical environments, the stakes around data fragmentation are highest. A multi-step QC workflow that spans HPLC, mass spectrometry, and a LIMS needs every handoff to be defensible during an audit, not just functional day to day.

The Pistoia Alliance’s MethodDB project, developed in collaboration with the Allotrope Foundation and partners including Agilent, Merck, and Bristol-Myers Squibb, uses the Allotrope Data Format to standardise how analytical method descriptions are recorded, increasing data integrity and scientific reproducibility. This kind of standardisation is precisely what reduces the change-management burden GxP labs face when adopting any new automation layer, the data structure underneath stays consistent even as the tools on top evolve.

Chemetrix application specialists work directly with regulated labs across the region to assess where data handoffs currently rely on manual reformatting, and to map a practical path toward standards-based interoperability without disrupting validated workflows already in place.


Improving cross-site asset utilisation for multi-lab operations

For organisations running multiple lab sites, instrument-level smart features solve a single-site problem while leaving a bigger one untouched: knowing which instruments across the network are actually available, and routing samples to them efficiently.

This is the orchestration layer gap that the broader industry has started naming directly. Coordination “still happens in calendars and email threads” in many R&D organisations, even where individual instruments are highly automated. Declarative orchestration, built on open data standards, is what allows a scheduling system to see instrument availability across sites and route work accordingly, rather than relying on a person manually checking which lab has capacity.

For multi-site operations evaluating this shift, Chemetrix can walk through a practical asset-utilisation audit, identifying where standards-based data exchange would unlock cross-site scheduling that today depends on manual coordination.

Should labs treat agentic AI as a cure-all for workflow problems?

No, and treating it that way is precisely the steep change-management trap that catches labs out. Agentic AI is only as effective as the data structure underneath it.

Agentic AI systems capable of autonomous planning, tool use, and multi-step reasoning represent a genuinely transformative shift, but the implications come with real governance challenges: how decisions are validated, how audit trails are maintained, and what infrastructure is needed to deploy agents safely at scale. A chatbot layered on top of fragmented, vendor-locked data doesn’t solve the underlying problem, it just adds a conversational interface to the same broken handoffs.

This is the cultural shift Chemetrix advocates for: building the data foundation first, with open standards and genuine interoperability, rather than reaching for an AI overlay as a shortcut around the harder infrastructure work. Labs that get this sequence right end up with systems that are auditable, defensible, and genuinely faster. Labs that skip ahead end up with an impressive demo and the same fragmented data problem underneath.

What should lab directors and IT architects do next?

The shift from instrument-centric benches to integrated, workflow-first ecosystems isn’t optional for labs that want to capture real efficiency gains, but it does require sequencing the work correctly: assess where data handoffs are currently manual, prioritise standards-based interoperability over point-solution AI overlays, and treat regulated change-management requirements as part of the plan from the outset, not an afterthought.

For Lab Directors: Save this guide and start mapping where your current workflow relies on manual data reformatting between instruments.
For IT Architects: Talk to a Chemetrix specialist about evaluating your lab’s readiness for open-standards-based data exchange.
For Operations Managers: Request an asset-utilisation audit to identify where declarative orchestration could improve cross-site scheduling and turnaround time.


TL;DR

First-generation “smart” lab features, automated peak integration, predictive maintenance alerts, solve problems inside a single instrument, but they don’t solve the data gaps between instruments. The real bottleneck is fragmented, vendor-locked data. Open standards like Allotrope, SiLA and the Pistoia Alliance’s frameworks are what let labs move from imperative integration (manual, rigid data mapping) to declarative orchestration, where an analyst sets a scientific goal and software coordinates the fleet, improving turnaround, compliance,
and asset utilisation.

How to Get Reliable FTIR Identification Every Time with the Agilent Cary 630

Your raw material just failed its FTIR identity check. The match score came back at 87%, the batch is on hold and three people are standing around a spectrometer debating whether to reject a container of perfectly good excipient.

Here is the hard truth: in most cases, the material is fine. The instrument is fine. The problem is a misunderstanding of what FTIR matching actually measures, and it is costing pharmaceutical QC labs hours of unnecessary retesting, false rejections and eroded confidence in a technology that is genuinely excellent at its job.

This article unpacks the three real causes of low match scores, what FTIR can and cannot tell you, and how to build a workflow that turns your spectrometer into the trusted identification workhorse it was designed to be.


Why does my FTIR keep giving me low match scores?

Walk into most pharmaceutical QC labs and you will see some version of the same frustration. A technician places a powder loosely on the diamond crystal, presses the plunger halfway down, gets a poor match score and logs it as a suspected material failure. After years of working alongside QC teams across hundreds of laboratories, the Chemetrix team can tell you that this scenario almost always comes down to one of three things.

Poor ATR crystal contact: Infrared light only penetrates a few micrometres into the sample surface. Air gaps or inconsistent pressure mean the evanescent wave never reaches the material properly, producing a weak and noisy spectrum that will not match any reference cleanly.

Generic reference libraries: Most labs rely on commercial libraries built from a single reference sample under ideal conditions. When your actual material arrives with slight batch variation, a different particle size or minor additives, it will not match that idealised reference, even if it is exactly the right material.

Misplaced expectations: A match score measures similarity under the conditions of measurement. It is not an absolute verdict on material identity. Even a correct material from the correct supplier can return a score well below 100% if technique or library quality is off.

The good news is that all three of these problems are fixable.

What is FTIR actually measuring?

FTIR spectroscopy works by focussing on infrared light at a sample and measuring which wavelengths the material absorbs. Every compound has a unique pattern of absorption peaks, its molecular fingerprint. The ATR (attenuated total reflectance) method makes this fast and practical by bouncing the infrared beam off a diamond crystal in contact with the sample, requiring no preparation beyond placing the material on the sensor.

Library matching then compares your sample’s fingerprint to a reference spectrum using a mathematical correlation algorithm. The result is a match score, not a binary yes or no,
but a measure of how similar the two spectra are under the conditions of that measurement. Importantly, standard library search methods are designed to detect differences in the 5 to 10% range between a sample and its reference, which means some variation is expected
and entirely normal.

This is a critical distinction. Natural spectral variation occurs between batches of the same material due to:

  • Moisture content and surface effects
  • Particle size differences between suppliers or manufacturing runs
  • Trace additives or stabilisers that are entirely acceptable within specification

None of these make the material wrong. But all of them will affect your match score if your reference library was not built to account for them.

Here is the insight that changes everything:

FTIR is an identification tool, not a purity tool. It will tell you what something is. It will not tell you how pure it is or quantify trace impurities. That is the job of HPLC. Stop asking your FTIR to do HPLC’s work and start letting it be the world-class ID workhorse it was built to be.

How do you improve ATR sample preparation?

Consistent results require consistent technique.
These are the four rules that make the biggest difference:

Powders: Spread the sample evenly across the full crystal surface before pressing. Apply firm and consistent downward pressure using the sample press. If the signal is weak, press harder and recollect the background.

Hard solids: Press a flat face directly onto the crystal or grind a small portion to a fine powder first. A smooth surface fragment beats a large irregular chunk every time.

Pastes and semi-solids: Apply a thin even layer onto the ATR surface with a spatula. These conform naturally to the crystal and are the easiest sample type to handle well.

Crystal hygiene: Clean the crystal between every sample with an appropriate solvent and a lint-free cloth. A contaminated crystal from the previous sample is an invisible source of error that will affect match scores across multiple runs.

Standardising these steps into a written SOP, what Chemetrix calls the “SOP of the Squeeze”, is the single fastest way to improve reproducibility across your team.

How can you build a pharmaceutical FTIR library that actually works?

The solution to false rejections in pharmaceutical raw material identification is not to lower your acceptance thresholds. It is to build a smarter library. A site-specific library collects reference spectra from your own approved batches of each material, across multiple suppliers and multiple delivery lots, so that the natural variation of your actual materials is built into the reference from the start. When a new batch arrives with slightly different moisture content or particle size, your library recognises it as the correct material because it has seen that variation before.

The instrument that makes this possible: the Agilent Cary 630 FTIR

The Cary 630 is purpose-built for pharmaceutical QC applications. Its instrument configuration delivers energy throughput up to 30% greater than comparable routine FTIR systems, which means faster data collection, lower noise and more reliable spectra even when sample variation is present. Its diamond ATR crystal is impervious to abrasion, requires very small amounts of sample and is easy to clean between samples. Modular sampling accessories click in and out in seconds with no alignment required, and at just 20 x 20 cm on the bench and 3.8 kg, it lives where the work happens rather than in a centralised lab.


Most importantly, the Cary 630 runs Agilent MicroLab software, a method-driven and pictorial interface that makes building site-specific libraries an intuitive task rather than a data science project. MicroLab supports advanced classification methods including PLS-DA (partial least squares discriminant analysis). In published testing using the Cary 630 with MicroLab, a PLS-DA method achieved 100% correct classification of pharmaceutical samples and detected contaminants at levels as low as 0.5%. Results are displayed as colour-coded pass/fail indicators and automatically logged in 21 CFR Part 11 compliant audit trails.

This is what Chemetrix calls the Golden Reference Library: a site-specific and validated collection of spectra that reflects your materials, your suppliers and your real-world conditions. Not a generic database. Yours.


Practical resources:

What does a validated FTIR library mean for regulatory compliance?

For pharmaceutical manufacturers operating in a regulated environment, a site-specific library is not just a practical improvement. It is a compliance asset. The Cary 630 meets or exceeds the performance specifications of the US, European, Japanese, Chinese, Indian and International Pharmacopoeia. Its automated IQ/OQ qualification software generates documented and auditable performance reports covering wavenumber accuracy, spectral resolution and signal-to-noise ratio. MicroLab OQ handles qualification for the US, European and Indian Pharmacopoeia directly, while MicroLab PC can be configured for additional requirements including the Japanese Pharmacopoeia. The optional MicroLab Pharma software package adds full 21 CFR Part 11 and EU Annex 11 support including electronic signatures and data integrity controls.


But instrument qualification is only one part of the compliance picture. A validated reference library, with documented collection conditions, approved batch traceability and defined acceptance criteria, is what gives your QA team the scientific and regulatory confidence to act on FTIR results without
second-guessing every match score.

Chemetrix supports clients through the full validation process, from initial library design through to IQ/OQ documentation and method verification against pharmacopoeial requirements.


Practical resources:

📌 Contact Chemetrix to discuss validation support for your FTIR workflow.

There is a better way to run your QC lab

Too many pharmaceutical QC labs have quietly accepted that FTIR is unreliable. They have added extra retesting steps. They have lowered thresholds without understanding why. They have learned to work around the instrument rather than with it. That is not good science. And it is not a sustainable way to run a quality control operation. Chemetrix believes that every lab, regardless of its size, its budget or its current frustrations, deserves instrumentation that works, workflows that are documented and a team that understands what their results actually mean.

The Cary 630 is not just sold and installed. Chemetrix partners with your team through three practical engagement points:

  • Workflow audit: Identifying exactly why your current match scores are low and where the process is breaking down
  • Method development Assistance: Building and documenting the SOP of the Squeeze that standardises ATR technique across your team
  • Validation support: Ensuring your custom library meets the compliance requirements of your QC environment

When your library is right and your technique is standardised, FTIR stops being a source of anxiety. It becomes what it was always supposed to be: fast, reliable and auditable identification, every time.


Low FTIR match scores are almost never an instrument problem.
They are a library problem or a technique problem, and both are fixable.

The three things to take away from this article:

    • A match score measures similarity under the conditions of measurement. Natural batch variation, particle size differences and trace additives will affect it even when the
      material is correct.
    • ATR sample preparation technique has a bigger impact on match scores than most labs realise. Standardising your approach with a documented SOP is the fastest win available.
    • A site-specific library built from your own approved batches will outperform any generic commercial database for your materials, in your lab and from your suppliers.

The Agilent Cary 630 FTIR makes the instrument side of this straightforward. Chemetrix makes the rest straightforward too.

Ready to stop second-guessing your FTIR results?

📩 Contact the Chemetrix team to book a workflow audit, arrange a Cary 630 demonstration or discuss building your Golden Reference Library: chemetrix.co.za

Powerful Partnerships, Inspiring Insights.

For over 45 years, Chemetrix has been a trusted partner to laboratories across Southern and East Africa, providing world-class analytical instrumentation and unwavering support.

As the sole authorised distributor of Agilent Technologies’ chromatography, mass spectrometry, and spectroscopy products in the region, we deliver cutting-edge solutions tailored to your laboratory’s unique needs.

Our team of highly qualified professionals—experts in the natural, life, and applied sciences—offer comprehensive support, from technical sales to application insights. We collaborate closely with our clients, ensuring that each solution aligns with their specific objectives.

Beyond our commitment to scientific excellence, Chemetrix is dedicated to social responsibility. Since 2017, we’ve partnered with eDeaf to provide skills development for deaf youth and actively participate in the Yes4Youth initiative, addressing youth unemployment in South Africa.

Whether you’re scaling up operations, navigating regulatory complexities, or seeking to enhance efficiency, Chemetrix stands ready to support your journey.

Let’s advance science together.
Connect with our team to discover how Chemetrix can elevate your laboratory’s performance.

AFSW2022: The Food Safety Event of the Year!

Africa Food Safety Workshop 2022 is happening at Emperors Palace in Johannesburg from 27 June until 1 July. The event brings together food safety stakeholders from across the continent to enhance networking and to facilitate the sharing of analytical knowledge and transfer of developed technologies that pertain to all areas of food safety control systems. 


The programme

The AFSW2022 programme boasts diverse discussion panels along with influential scientists and key role players delivering presentations that aim to boost systems and enhance trade across the sector. Q&A sessions give attendees the opportunity to engage with the speakers and get additional insights on the exciting range of topics. The event will also have a dedicated exhibition for various Food Safety instrumentation and equipment suppliers, like Chemetrix. 

You can view the full programme here


Why should you attend?

With the globalisation of the food chain, there is a greater demand for consistent quality, uncompromising safety and a commitment to protecting consumers while also adopting sustainable solutions. Chemetrix leads the industry with products and services to help you deliver what your customers demand. Our instruments, systems, and supplies are used throughout the food production chain, including incoming inspection, new product development, quality control and assurance, and packaging. We are excited to showcase our products at AFSW2022 along with our expertise, and we look forward to networking with the industry professionals in attendance. 

This event is important to members of various communities connected to food safety, including: 

  • Food testing laboratories (analysts and managers)
  • Quality infrastructure institutions including Accreditation, Standards, Scientific and Legal Metrology institutes
  • Regulators
  • Research/academia
  • Industry (food manufacturers/distributors)
  • International and non-governmental organizations
  • Professional associations


The event will be led by Keynote Speaker Dr. Godfrey Bahiigwa (African Union Commission – Director of the Department of Agriculture, Rural Development, Blue Economy and Sustainable Environment (DARBE), AU, Ethiopia).


For more event information, visit afsw2022.co.za to purchase your tickets and you can see the discussion and speaker programme here.